
9.6: Hückel MO Model of Conjugation
- Page ID
- 518983

\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\avec}{\mathbf a}\) \(\newcommand{\bvec}{\mathbf b}\) \(\newcommand{\cvec}{\mathbf c}\) \(\newcommand{\dvec}{\mathbf d}\) \(\newcommand{\dtil}{\widetilde{\mathbf d}}\) \(\newcommand{\evec}{\mathbf e}\) \(\newcommand{\fvec}{\mathbf f}\) \(\newcommand{\nvec}{\mathbf n}\) \(\newcommand{\pvec}{\mathbf p}\) \(\newcommand{\qvec}{\mathbf q}\) \(\newcommand{\svec}{\mathbf s}\) \(\newcommand{\tvec}{\mathbf t}\) \(\newcommand{\uvec}{\mathbf u}\) \(\newcommand{\vvec}{\mathbf v}\) \(\newcommand{\wvec}{\mathbf w}\) \(\newcommand{\xvec}{\mathbf x}\) \(\newcommand{\yvec}{\mathbf y}\) \(\newcommand{\zvec}{\mathbf z}\) \(\newcommand{\rvec}{\mathbf r}\) \(\newcommand{\mvec}{\mathbf m}\) \(\newcommand{\zerovec}{\mathbf 0}\) \(\newcommand{\onevec}{\mathbf 1}\) \(\newcommand{\real}{\mathbb R}\) \(\newcommand{\twovec}[2]{\left[\begin{array}{r}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\ctwovec}[2]{\left[\begin{array}{c}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\threevec}[3]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\cthreevec}[3]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\fourvec}[4]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\cfourvec}[4]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\fivevec}[5]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\cfivevec}[5]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\mattwo}[4]{\left[\begin{array}{rr}#1 \amp #2 \\ #3 \amp #4 \\ \end{array}\right]}\) \(\newcommand{\laspan}[1]{\text{Span}\{#1\}}\) \(\newcommand{\bcal}{\cal B}\) \(\newcommand{\ccal}{\cal C}\) \(\newcommand{\scal}{\cal S}\) \(\newcommand{\wcal}{\cal W}\) \(\newcommand{\ecal}{\cal E}\) \(\newcommand{\coords}[2]{\left\{#1\right\}_{#2}}\) \(\newcommand{\gray}[1]{\color{gray}{#1}}\) \(\newcommand{\lgray}[1]{\color{lightgray}{#1}}\) \(\newcommand{\rank}{\operatorname{rank}}\) \(\newcommand{\row}{\text{Row}}\) \(\newcommand{\col}{\text{Col}}\) \(\renewcommand{\row}{\text{Row}}\) \(\newcommand{\nul}{\text{Nul}}\) \(\newcommand{\var}{\text{Var}}\) \(\newcommand{\corr}{\text{corr}}\) \(\newcommand{\len}[1]{\left|#1\right|}\) \(\newcommand{\bbar}{\overline{\bvec}}\) \(\newcommand{\bhat}{\widehat{\bvec}}\) \(\newcommand{\bperp}{\bvec^\perp}\) \(\newcommand{\xhat}{\widehat{\xvec}}\) \(\newcommand{\vhat}{\widehat{\vvec}}\) \(\newcommand{\uhat}{\widehat{\uvec}}\) \(\newcommand{\what}{\widehat{\wvec}}\) \(\newcommand{\Sighat}{\widehat{\Sigma}}\) \(\newcommand{\lt}{<}\) \(\newcommand{\gt}{>}\) \(\newcommand{\amp}{&}\) \(\definecolor{fillinmathshade}{gray}{0.9}\)- Demonstrate how Hückel's theory approximates the full molecular orbital picture of molecules by treating the \(\sigma\)-bonding and \(\pi\)-bonding networks independently.
Introduction
Molecular orbital theory has been very successfully applied to large conjugated systems, especially those containing chains of carbon atoms with alternating single and double bonds. An approximation introduced by Hückel in 1931 considers only the delocalized p electrons moving in a framework of \(\pi\)-bonds. This is, in fact, a more sophisticated version of a free-electron model.
The simplest hydrocarbon to consider that exhibits \(\pi\) bonding is ethylene (ethene), which is made up of four hydrogen atoms and two carbon atoms. Experimentally, we know that the H–C–H and H–C–C angles in ethylene are approximately 120°. This angle suggests that the carbon atoms are sp2 hybridized, which means that a singly occupied sp2 orbital on one carbon overlaps with a singly occupied s orbital on each H and a singly occupied sp2 lobe on the other C. Thus each carbon forms a set of three \(\sigma\) bonds: two C–H (sp2 + s) and one C–C (sp2 + sp2) (part (a) of Figure \(\PageIndex{1}\) ).

The Hückel approximation is used to determine the energies and shapes of the \(\pi\) molecular orbitals in conjugated systems. Within the Hückel approximation, the covalent bonding in these hydrocarbons is separated into two independent "frameworks": the \(\sigma\)-bonding framework and the the \(\pi\)-bonding framework. Only \(\pi\) electron molecular orbitals are included because these determine the general properties of these molecules; the \(\sigma\) electrons are ignored. This is referred to as sigma-pi separability and is justified by the orthogonality of \(\sigma\) and \(\pi\) orbitals in planar molecules. For this reason, the Hückel method is limited to planar systems. The Hückel approximation focuses only on the formation of \(\pi\) bonds, given that the \(\sigma\) bonding framework has already been formed.
A conjugated system has a region of overlapping p-orbitals, bridging the interjacent single bonds, that allow a delocalization of \(\pi\) electrons across all the adjacent aligned p-orbitals. These \(\pi\) electrons do not belong to a single bond or atom, but rather to a group of atoms.
\(\pi\) system of ethylene
The \(\sigma\)-bonding occurs via the mixing of the electrons in the \(sp^2\) hybrid orbitals on carbon and the electrons in the \(1s\) atomic orbitals of the four hydrogen atoms (Figure \(\PageIndex{1}\) (a)). The \(\pi\)-bonding framework results from the unhybridized \(2p_z\) orbitals (Figure \(\PageIndex{1}\) (b)). The independence of these two frameworks is demonstrated in the resulting molecular orbital diagram in Figure \(\PageIndex{2}\).

In Hückel theory the molecular orbitals \(\psi_i \) can be described as a linear combination of the \(2p_z\) atomic orbitals \(\phi\) at each carbon with their corresponding \(\{c_i\}\) coefficients:
\[ \psi_i =c_1 \phi_{1} +c_2 \phi_2 \label{LCAO} \]
Substituting this into the Schrödinger equation:
\[ \hat{H} \psi_i =E_i \psi_i \nonumber \]
with \(\hat{H}\) the Hamiltonian and \(E_i\) the energy corresponding to the molecular orbital to give:
\[ \hat{H} c_{1} \phi _{1} +\hat{H} c_{2} \phi _{2} =Ec_{1} \phi _{1} +Ec_{2} \phi _{2} \label{SEq} \]
Notice that the E is for the hybrid orbital, so does not have a subcript. If Equation \(\ref{SEq}\) is left multiplied by \(\phi _{1}^* \) (and integrated), then
\[\int_{all space} \phi _{1}^* \hat{H} c_{1} \phi _{1} + \phi _{1}^* \hat{H} c_{2} \phi _{2} d\tau=\int_{all space}\phi _{1}^* E c_{1} \phi _{1} +\phi _{1}^*E c_{2} \phi _{2} d \tau \nonumber\]
Utilizing the fact that the c's and E are constants this rearranges to:
\[c_{1}\int_{all space} \phi _{1}^* \hat{H} \phi _{1} d\tau + c_{2}\int_{all space} \phi _{1}^* \hat{H} \phi _{2} d\tau=E c_{1}\int_{all space}\phi _{1}^* \phi _{1} d\tau +E c_{2}\int_{all space}\phi _{1}^* \phi _{2} d \tau \nonumber\]
Using Hik to represent the integrals with Hamiltonians in them and Sik to represent overlap integrals (no Hamiltonian between the wavefunctions) of orbital i and k this can be written more compactly as:
\[c_1H_{11} + c_2H_{12} = c_1 E S_{11} + c_2 E S_{12}\nonumber\]
collecting all the terms on one side yields the first of a system of equations in the coefficients:
\[c_1(H_{11} - E S_{11}) + c_2(H_{12} - E S_{12}) = 0 \label{Eq1} \]
The same analysis but starting by left multiplying by \(\phi_2^*\) and integrating over all of space leads to the second:
\[c_1(H_{21} - E S_{21}) + c_2(H_{22} - E S_{22}) = 0 \label{Eq2} \]
Nontrivial solutions, \(c_i \ne 0\), only exist for the coefficients if the determinant of the matrix of the integral quantities in parentheses (the secular determinant) is equal to zero.
\[ \left| \begin{array} {cc} H_{11} - ES_{11} & H_{12} - ES_{12} \\ H_{21} - ES_{21} & H_{22} - ES_{22} \\ \end{array}\right| = 0 \label{SecDet} \]
All diagonal Hamiltonian integrals \( H_{ii}\) are called Coulomb integrals and those of type \(H_{ik}\) are called resonance integrals. Both integrals are negative and the resonance integrals determines the strength of the bonding interactions. The Sik integrals are called the overlap integrals and Sii = 1 for normalized wavefunctions. The equations described in \(\ref{Eq1}\) and \(\ref{Eq2}\) are called the secular equations.
Everything in Equation \(\ref{SecDet}\) is a known number except \(E\). Since the secular determinant for ethylene is a \(2 \times 2\) matrix, finding \(E\), requires solving a quadratic equation (after expanding the determinant):
\[ ( H_{11} - ES_{11} ) ( H_{22} - ES_{22} ) - ( H_{21} - ES_{21} )( H_{12} - ES_{12} ) = 0\nonumber \]
There will be two values of \(E\) which satisfy this equation and they are the molecular orbital energies. For ethylene, one will be the bonding energy and the other the antibonding energy for the \(\pi\)-orbitals formed by the combination of the two carbon \(2p_z\) orbitals (Equation \(\ref{LCAO}\)). However, if more than two \(\phi\) atomic orbitals were used, e.g., in a bigger molecule, then more energies would be estimated by solving the secular determinant.
A 2 X 2 determinant is calculated as follows:
\[\begin{vmatrix} a &b \\ c &d \end{vmatrix} = ad - cb \]
A 3 X 3 determinant is calculated as follows:
\[\begin{vmatrix} a_{11} &a_{12} & a_{13} \\ a_{21} &a_{22} &a_{23} \\ a_{31} &a_{32} &a_{33} \end{vmatrix}= a_{11}\begin{vmatrix} a_{22}&a_{23}\\ a_{32}&a_{33} \end{vmatrix}- a_{12}\begin{vmatrix} a_{21}&a_{23}\\ a_{31}&a_{33} \end{vmatrix}+ a_{13}\begin{vmatrix} a_{21}&a_{22}\\ a_{31}&a_{32} \end{vmatrix}\]
This drawing indicates how you pick the sub-determinants:

For more on determinants see: https://chem.libretexts.org/Bookshel...A_Determinants .
Applying the Hückel approximations
Solving the secular determinant is simplified in the Hückel method via the following four assumptions:
- All overlap integrals \(S_{ik}\) with k ≠ i are set equal to zero. This is quite reasonable since the \(\pi-\) orbitals are directed perpendicular to the direction of their bonds (Figure \(\PageIndex{1}\) ). This assumption is often call neglect of differential overlap (NDO).
- All resonance integrals \(H_{ik}\) between non-neighboring atoms are set equal to zero.
- All resonance integrals \(H_{ik}\) between neighboring atoms are equal and set to \(\beta\).
- All coulomb integrals \(H_{ii}\) are set equal to \(\alpha\).
These assumptions are mathematically expressed as
\[ H_{11}=H_{22}=\alpha\nonumber \]
\[ H_{12}=H_{21}=\beta\nonumber \]
Assumptions 1 means that the overlap integral between any two atomic orbitals on different atoms is 0. Thus:
\[ S_{11}=S_{22}=1\nonumber \]
\[ S_{12}=S_{21}=0\nonumber \]
The Hückel assumptions reduces Equation \(\ref{SecDet}\) to:
\[\begin{vmatrix} \alpha - E & \beta \\ \beta & \alpha - E \\ \end{vmatrix}= 0 \label{Eq12} \]
if Equation \(\ref{Eq12}\) is divided by \(\beta\):
\[\begin{vmatrix} \dfrac{\alpha - E}{\beta} & 1 \\ 1 & \dfrac{\alpha - E}{\beta} \\ \end{vmatrix}= 0\label{eq13} \]
and then a new variable \(x\) is defined
\[ x = \dfrac {\alpha -E}{\beta} \label{new} \]
then Equation \(\ref{eq13}\) simplifies to
\[ \begin{vmatrix}x&1\\1&x\\\end{vmatrix}=0\nonumber \]
The trivial solution gives both wavefunction coefficients equal to zero and the other (nontrivial) solution is determined by solving the secular determinant
\[ \begin{vmatrix}x&1\\1&x\\\end{vmatrix} = (x-1)(x+1)=0\nonumber \]
so \( x=\pm 1\).
Knowing that \(E=\alpha -x\beta \) from Equation \(\ref{new}\), the energy levels can be found to be
\[ E=\alpha -\pm 1\times \beta \nonumber \]
or
\[ E=\alpha \mp \beta \nonumber \]
Since \(\beta\) is negative, the two energies are ordered (Figure \(\PageIndex{3}\))
- For \(\pi_1\): \(E_1 =\alpha + \beta\)
- For \(\pi_2\): \(E_2 =\alpha - \beta\)

To extract the coefficients attributed to these energies, the corresponding E values can be substituted back into the secular equations \(\ref{Eq1}\) and \(\ref{Eq2}\) and combined with the normalization requirement. For the lower energy state (\(E_1 =\alpha + \beta\))
\[c_1(H_{11} - E S_{11}) + c_2(H_{12} - E S_{12}) = c_1(\alpha - (\alpha + \beta) ) + c_2(\beta - (\alpha + \beta) (0)) = c_1(-\beta) ) + c_2\beta= 0 \label{Eq1b} \]
and
\[c_1(H_{21} - E S_{21}) + c_2(H_{22} - E S_{22}) = c_1(\beta - (\alpha + \beta) (0)) + c_2(\alpha - (\alpha + \beta)) =c_1\beta - c_2\beta = 0 \label{Eq2b} \]
This gives \(c_1=c_2\) and the molecular orbitals attributed to this energy is then (based off of Equation \(\ref{LCAO}\)):
\[ \psi_1 = N_1 (\phi_1 + \phi_2 ) \label{HOMO} \]
where \(N_1\) is the normalization constant for this molecular orbital; this is the bonding molecular orbital.
For the higher energy molecular substituting in the energy gives \(c_1=-c_2\) and the molecular orbitals attributed to this energy is then:
\[ \psi_2 = N_2 (\phi_1 - \phi_2 ) \label{LUMO} \]
where \(N_2\) is the normalization constant for this molecular orbital; this is the anti-bonding molecular orbital.
Normalization of \(\psi_1\), remembering the \(\phi\) are real and Sik = 0 when i ≠ k, leads to:
\[1 = \int_{\text{all space}}\psi_1^* \psi_1 d\tau = \int_{\text{all space}}N_1 (\phi_1 + \phi_2 )N_1 (\phi_1 + \phi_2 ) d\tau = N_1^2(S_{11} + S_{22}) = 2N_1^2\nonumber\]
and similarly for \(\psi_2\):
\[1 = 2N_2^2\nonumber\]
Thus:
\[N_1 = N_2 = \dfrac{1}{\sqrt{2}}\nonumber \]
These molecular orbitals form the \(\pi\)-bonding framework and since each carbon contributes one electron to this framework, only the lowest molecular orbital (\( \psi_1 \)) is occupied (Figure \(\PageIndex{4}\) ) in the ground state. The corresponding electron configuration is then \( \pi_1^2\).

HOMO and LUMO are acronyms for highest occupied molecular orbital and lowest unoccupied molecular orbital, respectively and are often referred to as frontier orbitals. The energy difference between the HOMO and LUMO is termed the HOMO–LUMO gap.
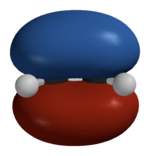
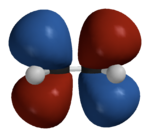
The 3-D calculated \(\pi\) molecular orbitals based on more sophisticated quantum calculations are shown in Figure \(\PageIndex{5}\).
Hückel theory was developed in the 1930's when computers were unavailable and a simple mathematical approaches were very important for understanding experiment. Although the assumptions in Hückel theory are drastic they enabled the early calculations of molecular orbitals to be performed with mechanical calculators or by hand. Hückel Theory can be extended to address other types of atoms in conjugated molecules (e.g., nitrogen and oxygen). Moreover, it can be extended to also treat \(\sigma\) orbitals and this "Extended Hückel Theory" is still used today. Despite the utility of Hückel Theory, it is highly qualitative and we should remember the limitations of Hückel Theory:
- Hückel Theory is very approximate
- Hückel Theory cannot calculate energies accurately (electron-electron repulsion is not calculated)
- Hückel Theory typically overestimates predicted dipole moments
Hückel Theory is best used to provide simplified models for understanding chemistry and for a detailed understanding modern ab initio molecular methods are required.
Hückel model applied to butadiene
Butadiene \(\ce{H2C=CH-CH=CH2}\), has two conjugated bonds. Thus four carbons are involved in the \(\pi\) system. By analogy with ethylene we can set up a secular matrix to solve, where we place x down the diagonal and 1 in the matrix elements representing neighboring p orbitals. The rows and columns of the determinant can be thought of as numbered by orbital positions (on atoms 1, 2, 3, or 4) as suggested in this table:
| Atom Numbers | 1 | 2 | 3 | 4 |
| 1 | x | 1 | 0 | 0 |
| 2 | 1 | x | 1 | 0 |
| 3 | 0 | 1 | x | 1 |
| 4 | 0 | 0 | 1 | x |
This leads to the determinant:
\[\left|\begin{array}{cccc}x&1&0&0\\1&x&1&0\\0&1&x&1\\0&0&1&x\end{array}\right|=0\label{26} \]
This is essentially the connection matrix for the butadiene molecule. Each pair of connected atoms is represented by 1, each non-connected pair by 0 and each diagonal element by \(x\). Expansion of the determinant in Equation \(\ref{26}\) gives the 4th order polynomial equation
\[x^{4}-3x^{2}+1=0 \label{27} \]
While solving 4th order equations typically require numerical estimation, Equation \(\ref{27}\) can be further simplified by recognizing that it is a quadratic equation in terms of \(x^{2}\). Therefore, the roots are
\[x^{2}= \dfrac{3\pm\sqrt{5}}{2} \nonumber \]
or \(x=\pm\; 0.618\) and \(x= \pm\; 1.618\). Since \(\alpha\) and \(\beta\) are negative, these molecular orbital energies can ordered in terms of energy (from lowest to highest):
\[E_1=\alpha+1.618\beta \label{E1} \]
\[E_2=\alpha+0.618\beta \label{E2} \]
\[E_3=\alpha-0.618\beta \label{E3} \]
\[E_4=\alpha-1.618\beta \label{E4} \]
This sequence of energies is displayed in the energy diagram of Figure \(\PageIndex{6}\).
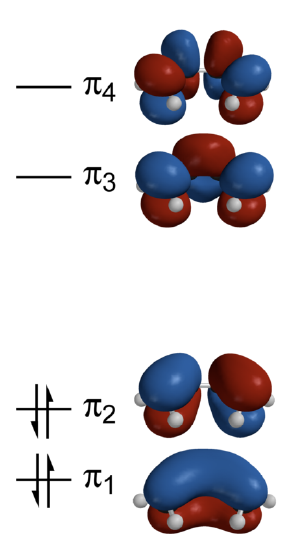
Each p atomic orbital of carbon contributes a single electron to the \(\pi\) manifold, so the ground-state occupation of the resulting four \(\pi\) electrons have a \(\pi_1^{2}\pi_2^{2}\) configuration (Figure \(\PageIndex{6}\) ). The the total \(\pi\)-electron energy is then determined by adding up the energies in Equations \(\ref{E1}\)-\(\ref{E4}\) and scaling by their occupations to get
\[\begin{align} E_{\pi} (\text{butadiene}) &= 2 \times E_1 + 2 \times E_2 + 0 \times E_3 + 0 \times E_4 \nonumber \\[4pt] &=2(\alpha+1.618\beta)+2(\alpha+0.618\beta) \nonumber \\[4pt] &=4\alpha + 4.472\beta \label{29} \end{align} \]
If the bonding of butadiene were described only as two localized double bond as in its dominant valence-bond structure, then its \(\pi\)-electron energy would be given by twice the \(E_{\pi}\) predicted for the ethlyene molecule:
\[\begin{align} E_{\pi} (\text{butadiene}) &= 2 \times E_{\pi} (\text{ethylene}) \nonumber \\[4pt] &=2 \times 2(\alpha+\beta) \nonumber \\[4pt] &= 4\alpha + 4\beta \label{30}\end{align} \]
Comparing Equation \(\ref{29}\) with Equation \(\ref{30}\), the total \(\pi\) energy of butadiene lies lower than the total \(\pi\) energy of two double bonds by \(0.472\beta\) (the \(\sigma\) bond does not contribute). This difference is known as the delocalization energy; a typical estimate of \(\beta\) is around -75 kJ/mol, which results in a delocalization energy for butadiene of -35 kJ/mol.
The delocalization energy is the extra stabilization resulting from the electrons extending over the whole molecule.
Delocalization energy is intrinsic to molecular orbital theory, since it results from breaking the two-center bond concept with the molecular orbitals that spread over more that just one pair of atoms. However, within the two-center theory of valence bond theory, the delocalization energy results from a stabilization energy attributed to resonance. Several conventional valence bond resonance structures can be written for 1,3-butadiene, four of which are shown in Figure \(\PageIndex{7}\). However, while the top left structure dominates, the other resonance structures also contribute to describing the total molecule and hence predict a corresponding stabilization energy akin to the delocalization energy in molecular orbital theory.

In general, the true description of the bonding within the valence bond theory is a superposition of resonance structures with amplitudes that are determined via a variational optimization to find the lowest possible energy for the valence bond wavefunctions.
Solving the secular equations (not shown) gives the weighting coefficients for the individual p orbital contributions to the molecular orbitals:
\[\psi_1 =0.37 p_1 + 0.60 p_2 + 0.60 p_3 + 0.37 p_4 \label{MO1} \]
\[\psi_3 =0.60 p_1 + 0.37 p_2 -0.37 p_3 - 0.60 p_4 \label{MO2} \]
\[\psi_3 =0.60 p_1 - 0.37 p_2 -0.37 p_3 + 0.60 p_4 \label{MO3} \]
\[\psi_4 =0.37 p_1 - 0.60 p_2 + 0.60 p_3 - 0.37 p_4 \label{MO4} \]
These are depicted in Figure \(\PageIndex{8}\) and using ab intio methods in Figure \(\PageIndex{6}\).

Note the correlation of the energy of the \(\pi\) molecular orbitals of butadiene to the number of nodes in the wavefunction; this is the general trend observed in previous systems like the particle in the box and atomic orbitals.
Contributors
-
Seymour Blinder (Professor Emeritus of Chemistry and Physics at the University of Michigan, Ann Arbor)
-
Delmar Larsen (UC Davis)
- Wikipedia
- StackExchange (Philipp)
- Jonathan Gutow (UW Oshkosh)