
5.2: Relaxation Methods
- Page ID
- 516736

\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\avec}{\mathbf a}\) \(\newcommand{\bvec}{\mathbf b}\) \(\newcommand{\cvec}{\mathbf c}\) \(\newcommand{\dvec}{\mathbf d}\) \(\newcommand{\dtil}{\widetilde{\mathbf d}}\) \(\newcommand{\evec}{\mathbf e}\) \(\newcommand{\fvec}{\mathbf f}\) \(\newcommand{\nvec}{\mathbf n}\) \(\newcommand{\pvec}{\mathbf p}\) \(\newcommand{\qvec}{\mathbf q}\) \(\newcommand{\svec}{\mathbf s}\) \(\newcommand{\tvec}{\mathbf t}\) \(\newcommand{\uvec}{\mathbf u}\) \(\newcommand{\vvec}{\mathbf v}\) \(\newcommand{\wvec}{\mathbf w}\) \(\newcommand{\xvec}{\mathbf x}\) \(\newcommand{\yvec}{\mathbf y}\) \(\newcommand{\zvec}{\mathbf z}\) \(\newcommand{\rvec}{\mathbf r}\) \(\newcommand{\mvec}{\mathbf m}\) \(\newcommand{\zerovec}{\mathbf 0}\) \(\newcommand{\onevec}{\mathbf 1}\) \(\newcommand{\real}{\mathbb R}\) \(\newcommand{\twovec}[2]{\left[\begin{array}{r}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\ctwovec}[2]{\left[\begin{array}{c}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\threevec}[3]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\cthreevec}[3]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\fourvec}[4]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\cfourvec}[4]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\fivevec}[5]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\cfivevec}[5]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\mattwo}[4]{\left[\begin{array}{rr}#1 \amp #2 \\ #3 \amp #4 \\ \end{array}\right]}\) \(\newcommand{\laspan}[1]{\text{Span}\{#1\}}\) \(\newcommand{\bcal}{\cal B}\) \(\newcommand{\ccal}{\cal C}\) \(\newcommand{\scal}{\cal S}\) \(\newcommand{\wcal}{\cal W}\) \(\newcommand{\ecal}{\cal E}\) \(\newcommand{\coords}[2]{\left\{#1\right\}_{#2}}\) \(\newcommand{\gray}[1]{\color{gray}{#1}}\) \(\newcommand{\lgray}[1]{\color{lightgray}{#1}}\) \(\newcommand{\rank}{\operatorname{rank}}\) \(\newcommand{\row}{\text{Row}}\) \(\newcommand{\col}{\text{Col}}\) \(\renewcommand{\row}{\text{Row}}\) \(\newcommand{\nul}{\text{Nul}}\) \(\newcommand{\var}{\text{Var}}\) \(\newcommand{\corr}{\text{corr}}\) \(\newcommand{\len}[1]{\left|#1\right|}\) \(\newcommand{\bbar}{\overline{\bvec}}\) \(\newcommand{\bhat}{\widehat{\bvec}}\) \(\newcommand{\bperp}{\bvec^\perp}\) \(\newcommand{\xhat}{\widehat{\xvec}}\) \(\newcommand{\vhat}{\widehat{\vvec}}\) \(\newcommand{\uhat}{\widehat{\uvec}}\) \(\newcommand{\what}{\widehat{\wvec}}\) \(\newcommand{\Sighat}{\widehat{\Sigma}}\) \(\newcommand{\lt}{<}\) \(\newcommand{\gt}{>}\) \(\newcommand{\amp}{&}\) \(\definecolor{fillinmathshade}{gray}{0.9}\)Explanation and Theory
Many reactions, especially those that take place in solution, occur too rapidly to follow by flow techniques, and can therefore only be observed when they are already at equilibrium. The classical examples of such reactions are two of the fastest ones ever observed, the dissociation of water
\[ 2 H_2O \rightarrow H_3O^+ + OH^- \nonumber \]
and the formation of the triiodide ion in aqueous solution
\[ I^– + I_2 \rightarrow I_3^– \nonumber \]
Reactions of these kinds could not be studied until the mid-1950s when techniques were developed to shift the equilibrium by imposing an abrupt physical change on the system.
For example, if the reaction A  B is endothermic, then according to the Le Chatelier principle, subjecting the system to a rapid jump in temperature will shift the equilibrium state to one in which the product B has a higher concentration. The composition of the system will than begin to shift toward the new equilibrium composition at a rate determined by the kinetics of the process. For this simple equilibrium, which is unimolecular in both directions, this can easily be interpreted in terms of a time dependent quantity xt = the distance to the new equilibrium (at t = 0, |xt| is at its maximum). Representing the time dependent concentrations as [A]t and [B]t while using [A]e and [B]e as the new equilibrium concentrations we can write the time dependent concentrations in terms of xt.
\[[A]_t = [A]_e + x_t\label{At}\]
\[[B]_t = [B]_e - x_t\label{Bt}\]
The rate laws for this equilibrium are:
\[\frac{d[A]_t}{dt} = -k_1[A]_t + k_{-1}[B]_t\label{dAdT}\]
\[\frac{d[B]_t}{dt} = k_1[A]_t - k_{-1}[B]_t\label{dBdT}\]
or
\[\text{rate} = -\frac{d[A]_t}{dt} =\frac{d[B]_t}{dt} = k_1[A]_t - k_{-1}[B]_t \label{rate}\]
Substituting into equation \(\ref{rate}\) the expressions for [A]t and [B]t (equations \(\ref{At}\) and \(\ref{Bt}\)) in terms of xt yields:
\[\text{rate} = -\frac{d([A]_e +x_t)}{dt} = k_1([A]_e +x_t) - k_{-1}([B]_e -x_t) \label{rate2}\]
Recognizing that \(\frac{d[A]_e}{dt} = 0\) because [A]e is a constant simplifies this to:
\[\text{rate} = -\frac{d x_t}{dt} = k_1[A]_e + k_1 x_t - k_{-1}[B]_e + k_{-1} x_t\label{rate3}\]
For any reaction at equilibrium the rate equals 0. Using the first rate law (equation \(\ref{dAdT}\)) as an example:
\[\frac{d[A]_t}{dt} = 0 = -k_1[A]_e + k_{-1}[B]_e\label{dAdTeq}\]
which can be rearranged to:
\[k_{-1}[B]_e = k_1[A]_e \label{ABrel}\]
or
\[K_{eq} = \frac{[B]_e}{[A]_e} = \frac{k_1}{k_{-1}} \label{Kc}\]
the concentration equilibrium constant. This relation between the concentration equilibrium constant and the ratios of the forward and reverse reactions is generally true for all reactions.
\(k_{-1}[B]_e = k_1[A]_e\) (equation \(\ref{ABrel}\)) can be used to replace \(k_{-1}[B]_e\) in equation \(\ref{rate3}\) yielding:
\[-\frac{d x_t}{dt} = k_1[A]_e + k_1 x_t - k_1[A]_e + k_{-1} x_t\label{rate4}\]
which simplifies to:
\[-\frac{d x_t}{dt} = k_1 x_t + k_{-1} x_t \implies \frac{d x_t}{dt} = -( k_1 + k_{-1}) x_t \label{rate5}\]
We recognize this as a first order process with the solution:
\[x_t = x_o exp \left(-(k_1 + k_{-1})t\right) = x_o exp \left(\frac{-t}{\tau}\right)\label{rate6}\]
where \(\tau = \frac{1}{k_1 + k_{-1}}\).
Although this derivation is for the unimolecular case, it can be shown that as long as xo is small first order kinetics will be observed for reactions of other molecularities (see for example: 2.1.4: Relaxation Methods, but be aware there are many typos despite the correct conclusions). The cartoon below illustrates what data may look like.
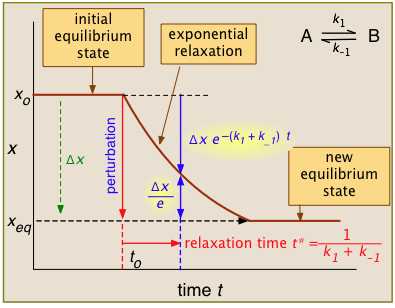
Fitting the exponential decay directly yields \(\tau\). The measurements also give us Keq (equation \(\ref{Kc}\), the ratio of the two rate constants). This gives us two equations in two unknowns. It is left to the reader to show that these can be solved for the two rate constants yielding:
\[k_1 = \frac{K_{eq}}{(1 + K_{eq})\tau} \quad\quad k_{-1} = \frac{1}{(1 + K_{eq})\tau} \nonumber\]
Similar analyses for higher order reactions yield somewhat more complicated expressions.
Examples
Temperature jumps
This is the method that Manfred Eigen (Germany, 1927-) pioneered when, in the early 1960's, he measured the rate constant of what was then the fastest reaction ever observed:
\[ H^+ + OH^– \rightarrow H_2O \;\;\;\; k = 1.3 \times 10^{11} \; M^{–1}\; sec^{–1} \nonumber \]
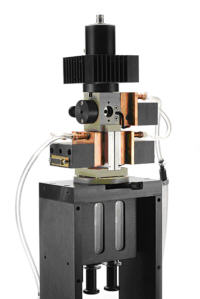
BioLogic rapid-mixing T-Jump System
Some ways of acheiving temperature jumps:
- high-voltage electric discharge: A capacitor, charged to 5-10 kV, is discharged through a solution to which an electrolyte has been added to provide a conductive path.
- laser irradiation: The sample is irradiated with a laser whose wavelength corresponds to an absorption peak in the sample. Infrared lasers are often used for this purpose.
- mixing of two pre-equilibrated solutions: Two solutions, otherwise identical but at different temperatures, are rapidly mixed in a stopped-flow type of apparatus. Although this method is not as fast, it has the advantage of allowing both negative and positive T-jumps. The device shown here uses 0.1-mL samples and provides jumps of up to ±40 C° over a few microseconds. Observation times, however, are limited to 1-2 milliseconds owing to thermal dissipation.
Pressure jumps
According to the Le Chatelier principle, a change in the applied pressure will shift the equilibrium state of any reaction which involves a change in the volume of a system. Aside from the obvious examples associated with changes in the number of moles of gases, there are many more subtle cases involving formation of complexes, hydration shells and surface adsorption, and phase changes. One area of considerable interest is the study of protein folding, which has implications in diseases such as Parkinson's and Alzheimer's.
The pressure-jump is applied to the cell through a flexible membrane that is activated by a high-pressure gas supply, or through an electrically-actuated piezoelectric crystal. This can produce P-jumps of around 1 GPa over sub-millisecond time intervals.
Shock tubes: extreme jumping
When a change in pressure propagates through a gas at a rate greater than the ordinary compressions and rarefactions associated with the travel of sound, a moving front (a shock wave) of very high pressure forms. This in turn generates an almost instantaneous rise in the temperature that can approach several thousand degrees in magnitude.
A shock tube is an apparatus in which shock waves can be generated and used to study the kinetics of gas-phase reactions that are otherwise inaccessible to kinetic measurements. Since all molecules tend to dissociate at high temperatures, shock tubes are widely used to study dissociation processes and the chemistry of the resulting fragments. For example, the shock-induced decomposition of carbon suboxide provides an efficient means of investigating carbon atom reactions:
\[ C_3O_2 \rightarrow C + CO \nonumber \]
Shock tube techniques are also useful for studying combustion reactions, including those that proceed explosively.
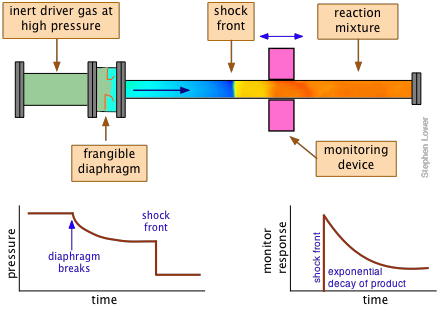
The shock tube itself consists of two sections separated by a breakable diaphragm of metal or plastic. One section is filled with a "driver" gas at a very high pressure, commonly helium, but often mixed with other inert gases to adjust the properties of the shock. The other, longer section of the tube contains the "driven" gas—the reactants—at a low pressure, usually less than 1 atmosphere.
The reaction is initiated by causing the diaphragm to rupture, either by means of a mechanical plunger or by raising the pressure beyond its bursting point. The kinetics of the reaction are monitored by means of an absorption or other optical monitoring device that is positioned at a location along the reaction tube that is appropriate to the time course of the reaction.
Contributors and Attributions
Stephen Lower, Professor Emeritus (Simon Fraser U.) Chem1 Virtual Textbook
- Jonathan Gutow (UW Oshkosh)