
1.8: Colligative Properties
- Page ID
- 498601

\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\avec}{\mathbf a}\) \(\newcommand{\bvec}{\mathbf b}\) \(\newcommand{\cvec}{\mathbf c}\) \(\newcommand{\dvec}{\mathbf d}\) \(\newcommand{\dtil}{\widetilde{\mathbf d}}\) \(\newcommand{\evec}{\mathbf e}\) \(\newcommand{\fvec}{\mathbf f}\) \(\newcommand{\nvec}{\mathbf n}\) \(\newcommand{\pvec}{\mathbf p}\) \(\newcommand{\qvec}{\mathbf q}\) \(\newcommand{\svec}{\mathbf s}\) \(\newcommand{\tvec}{\mathbf t}\) \(\newcommand{\uvec}{\mathbf u}\) \(\newcommand{\vvec}{\mathbf v}\) \(\newcommand{\wvec}{\mathbf w}\) \(\newcommand{\xvec}{\mathbf x}\) \(\newcommand{\yvec}{\mathbf y}\) \(\newcommand{\zvec}{\mathbf z}\) \(\newcommand{\rvec}{\mathbf r}\) \(\newcommand{\mvec}{\mathbf m}\) \(\newcommand{\zerovec}{\mathbf 0}\) \(\newcommand{\onevec}{\mathbf 1}\) \(\newcommand{\real}{\mathbb R}\) \(\newcommand{\twovec}[2]{\left[\begin{array}{r}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\ctwovec}[2]{\left[\begin{array}{c}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\threevec}[3]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\cthreevec}[3]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\fourvec}[4]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\cfourvec}[4]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\fivevec}[5]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\cfivevec}[5]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\mattwo}[4]{\left[\begin{array}{rr}#1 \amp #2 \\ #3 \amp #4 \\ \end{array}\right]}\) \(\newcommand{\laspan}[1]{\text{Span}\{#1\}}\) \(\newcommand{\bcal}{\cal B}\) \(\newcommand{\ccal}{\cal C}\) \(\newcommand{\scal}{\cal S}\) \(\newcommand{\wcal}{\cal W}\) \(\newcommand{\ecal}{\cal E}\) \(\newcommand{\coords}[2]{\left\{#1\right\}_{#2}}\) \(\newcommand{\gray}[1]{\color{gray}{#1}}\) \(\newcommand{\lgray}[1]{\color{lightgray}{#1}}\) \(\newcommand{\rank}{\operatorname{rank}}\) \(\newcommand{\row}{\text{Row}}\) \(\newcommand{\col}{\text{Col}}\) \(\renewcommand{\row}{\text{Row}}\) \(\newcommand{\nul}{\text{Nul}}\) \(\newcommand{\var}{\text{Var}}\) \(\newcommand{\corr}{\text{corr}}\) \(\newcommand{\len}[1]{\left|#1\right|}\) \(\newcommand{\bbar}{\overline{\bvec}}\) \(\newcommand{\bhat}{\widehat{\bvec}}\) \(\newcommand{\bperp}{\bvec^\perp}\) \(\newcommand{\xhat}{\widehat{\xvec}}\) \(\newcommand{\vhat}{\widehat{\vvec}}\) \(\newcommand{\uhat}{\widehat{\uvec}}\) \(\newcommand{\what}{\widehat{\wvec}}\) \(\newcommand{\Sighat}{\widehat{\Sigma}}\) \(\newcommand{\lt}{<}\) \(\newcommand{\gt}{>}\) \(\newcommand{\amp}{&}\) \(\definecolor{fillinmathshade}{gray}{0.9}\)Colligative properties are important properties of solutions as they describe how the properties of the solvent will change as solute (or solutes) is (are) added. Before discussing these important properties, let us first review some definitions.
- Solution – a homogeneous mixture.
- Solvent – The component of a solution with the largest mole fraction
- Solute – Any component of a solution that is not the solvent.
Solutions can exist in solid (alloys of metals are an example of solid-phase solutions), liquid, or gaseous forms. For the most part, this discussion will focus on liquid-phase solutions.
Freezing Point Depression
Empirically it is found that the change in freezing point is proportional to the molality of the solute:
\[\Delta T_f = K_f m_B\label{EQ:Fpdepr}\]
where \(\Delta T\) gives the magnitude of the reduction of freezing point for the solution, Kf is is the cryoscopic constant for the solvent and mB is the molality of the solute.
mB is the molality of particles dissolved not molality of empirical formula units. If the solute is NaCl than mB ≈ 2(molality of NaCl). This is often indicated by including the van 't Hoff factor (usually represented by i): \(\Delta T_f = K_f i m_B\). Because many salts do not dissociate completely except at very low concentrations the van 't Hoff factor is an additional empirically determined quantity.
In equation \(\ref{EQ:Fpdepr}\) Kf is a negative value. However, many tabulations record Kf as a positive value. You must remember that \(\Delta T\) for freezing point depression is < 0 and change the sign of tabulated Kf values accordingly.
Notice that this result is independent of the identity of the solute.
The solute particles do not dissolve in the solid. This means that adding solute to the liquid phase increases its entropy relative to the solid phase, favoring the liquid phase. Thus, the temperature must drop to a lower value to favor the solid phase. It is important to keep in mind that for a real solution, freezing of the solvent changes the composition of the solution by decreasing the mole fraction of the solvent and increasing that of the solute. As such, the magnitude of \(\Delta T\) will change as the freezing process continually removes solvent from the liquid phase of the solution.
Demonstration of why the dependence on concentration is expected to be nearly linear.
In general a liquid will freeze when
\[ \mu_{s} \le \mu_{l} \nonumber \]
The key is that the solute remains in the liquid solvent. As such, the freezing point of the solvent in a solution will be affected by anything that changes the chemical potential of the solvent, while the chemical potential of the solid will only be impacted by the temperature change. In a mixture, the chemical potential of component \(A\) can be calculated by
\[\mu_A=\mu_A^o + RT \ln x_A \label{EQ:chempsolvent} \]
And because \(x_A\) is always less than (or equal to) 1, the chemical potential is always reduced by the addition of another component.
We also know how temperature impacts chemical potential, which will effect both phases:
\[\left(\frac{\partial{\mu}}{\partial{T}}\right)_P = -S_{molar} \implies \Delta \mu = -S_{molar} \Delta T \label{EQ:chempDeltaT}\]
The condition under which the solvent will freeze is
\[ \mu_{s} = \mu_{l} \nonumber \]
If we start at the standard freezing point of the pure solvent we can call the chemical potential \(\mu^{fp}\). For the solid phase when we reach the new equilibrium this is only impacted by the temperature change, while the solvent chemical potential is changed by both dissolving the solute and the temperature change. Thus, at the new equilibrium we have new chemical potentials for the two phases:
\[\mu_s(T_2) = \mu^{fp} - S_{molar}(s)\Delta T \label{EQ:musolid}\]
\[\mu_l(T_2) = \mu^{fp} - S_{molar}(l)\Delta T + RT_2 ln(x_A)\label{EQ:muliquid}\]
Setting these equal to each other:
\[ \cancel{\mu^{fp}} - S_{molar}(s)\Delta T =\cancel{ \mu^{fp}} - S_{molar}(l)\Delta T + RT_2 ln(x_A) \implies - S_{molar}(s)\Delta T = - S_{molar}(l)\Delta T + RT_2 ln(x_A) \label{EQ:muT2}\]
Collecting the entropy terms on one side and recognizing that \(x_A = 1 - x_B\) yields:
\[ S_{molar}(l)\Delta T - S_{molar}(s)\Delta T = RT_2 ln(x_A) \implies \Delta S_{fus}\Delta T = RT_2 ln(1 - x_B)\label{EQ:muT3}\]
For small \(x_B\), \(ln(1-x_B) \approx -x_B\):
\[\Delta S_{fus}\Delta T = -RT_2 x_B\label{EQ:muT4}\]
which can be rearranged to:
\[\Delta T = -\frac{RT_2}{\Delta S_{fus}} x_B\label{EQ:cryoscopic}\]
Since we have assumed a relatively small \(x_B\) the temperature change will be small meaning \(\Delta S\) will be nearly constant. Thus, everything in the fraction on the right-hand-side of equation \(\ref{EQ:cryoscopic}\) can be identified as the constant \(K_f\). As molality is proportional to mole fraction this justifies the form of the freezing point depression equation.
Boiling Point Elevation
Boiling point elevation is similar to freezing point depression. In short, the introduction of a solute into a liquid solvent lowers the chemical potential of the solvent, causing it to favor the liquid phase over the vapor phase. As such, the temperature must be increased to increase the chemical potential of the solvent in the liquid solution until it is equal to that of the vapor-phase solvent. The increase in the boiling point can be expressed as
\[ \Delta T = K_b m_B \nonumber \]
\(K_b\) is called the ebullioscopic constant and, like the cryoscopic constant, is a property of the solvent that is independent of the solute or solutes.
As with freezing point depression \(m_B\) is the molality of dissolved particles.
Some values of \(K_f\) and \(K_b\) are shown in the table below.
| Substance | \(K_f\) (°C kg mol-1) | \(T^o_f\) (°C) | \(K_b\) (°C kg mol-1) | \(T^o_b\) (°C) |
|---|---|---|---|---|
| Water | 1.86 | 0.0 | 0.51 | 100.0 |
| Benzene | 5.12 | 5.5 | 2.53 | 80.1 |
| Ethanol | 1.99 | -114.6 | 1.22 | 78.4 |
| CCl4 | 29.8 | -22.3 | 5.02 | 76.8 |
The boiling point of a solution of 3.00 g of an unknown compound in 25.0 g of CCl4 raises the boiling point to 81.5 °C. What is the molar mass of the compound?
Solution
The approach here is to find the number of moles of solute in the solution. First, find the concentration of the solution:
\[(85.5\, °C- 76.8\, °C) = \left( 5.02\, °C\,Kg/mol \right) m \nonumber \]
\[ m= 0.936\, mol/kg \nonumber \]
Using the number of kg of solvent, one finds the number for moles of solute:
\[ \left( 0.936 \,mol/\cancel{kg} \right) (0.02\,\cancel{kg}) =0.0234 \, mol \nonumber \]
The ratio of mass to moles yields the final answer:
\[\dfrac{3.00 \,g}{0.0234} = 128 g/mol \nonumber \]
Vapor Pressure Lowering
For volatile solutes we have noted that Raoult's Law \(P_A = x_A P_A^*\) is usually only valid at low solute concentrations. The derivation below shows that for non-volatile solutes Raoult's law can be expected to work over a much larger mole fraction range.
For much the same reason as the lowering of freezing points and the elevation of boiling points for solvents into which a solute has been introduced, the vapor pressure of a volatile solvent will be decreased due to the introduction of a solute. The magnitude of this decrease can be quantified by examining the effect the solute has on the chemical potential of the solvent.
In order to establish equilibrium between the solvent in the solution and the solvent in the vapor phase above the solution, the chemical potentials of the two phases must be equal.
\[\mu_{vapor} = \mu_{solvent} \nonumber \]
If the solute is not volatile, the vapor will be pure, so (assuming ideal behavior)
\[\mu_{vap}^o + RT \ln \dfrac{P'}{P^o} = \mu_A^o + RT \ln x_A \label{eq3} \]
Where \(P’\) is the vapor pressure of the solvent over the solution. Similarly, for the pure solvent in equilibrium with its vapor
\[ \mu_A^o = \mu_{vap}^o + RT \ln \dfrac{P^*_A}{P^o} \label{eq4} \]
where \(P^o\) is the standard pressure of 1 atm, and \(P^*_A\) is the vapor pressure of the pure solvent. Substituting Equation \ref{eq4} into Equation \ref{eq3} yields
\[ \cancel{\mu_{vap}^o} + RT \ln \dfrac{P'}{P^o}= \left ( \cancel{\mu_{vap}^o} + RT \ln \dfrac{P^*_A}{P^o} \right) + RT \ln x_A \nonumber \]
The terms for \(\mu_{vap}^o\) cancel, leaving
\[ RT \ln \dfrac{P'}{P^o}= RT \ln \dfrac{P^*_A}{P^o} + RT \ln x_A \nonumber \]
Subtracting \(RT \ln(P^*_A/P^o)\) from both side produces
\[ RT \ln \dfrac{P'}{P^o} - RT \ln \dfrac{P^*_A}{P^o} = RT \ln x_A \nonumber \]
which rearranges to
\[RT \ln \dfrac{P'}{P^*_A} = RT \ln x_A \nonumber \]
Dividing both sides by \(RT\) and then exponentiating yields
\[ \dfrac{P'}{P^*_A} = x_A \nonumber \]
or
\[ P'=x_A P^*_A \label{RL} \]
This last result is Raoult’s Law. A more formal derivation would use the fugacities of the vapor phases, but would look essentially the same. Also, as in the case of freezing point depression and boiling point elevations, this derivation did not rely on the nature of the solute! However, unlike freezing point depression and boiling point elevation, this derivation did not rely on the solute being dilute, so the result should apply the entire range of concentrations of the solution.
Consider a mixture of two volatile liquids A and B. The vapor pressure of pure A is 150 Torr at some temperature, and that of pure B is 300 Torr at the same temperature. What is the total vapor pressure above a mixture of these compounds with the mole fraction of B of 0.600. What is the mole fraction of B in the vapor that is in equilibrium with the liquid mixture?
Solution
Using Raoult’s Law (Equation \ref{RL})
\[ P_A = (0.400)(150\, Toor) =60.0 \,Torr \nonumber \]
\[ P_B = (0.600)(300\, Toor) =180.0 \,Torr \nonumber \]
\[ P_{tot} = P_A + P_B = 240 \,Torr \nonumber \]
To get the mole fractions in the gas phase, one can use Dalton’s Law of partial pressures.
\[ x_A = \dfrac{ P_A}{P_{tot}} = \dfrac{60.0 \,Torr}{240\,Torr} = 0.250 \nonumber \]
\[ x_B = \dfrac{ P_B}{P_{tot}} = \dfrac{180.0 \,Torr}{240\,Torr} = 0.750 \nonumber \]
And, of course, it is also useful to note that the sum of the mole fractions is 1 (as it must be!)
\[ x_A+x_B =1 \nonumber \]
Osmotic Pressure and Osmotic Virial Coefficients
In dealing with solutions (either solutions of small molecule solutes or macromolecules in solution), an important colligative property is the osmotic pressure. The osmotic pressure is given by the symbol \( \Pi \) and is different from the pressure of a gas (P) because it does not arise from collisions of molecules against the wall of a container. The osmotic pressure is a hydrostatic pressure that arises when solvent molecules pass through a semipermeable membrane to the more concentrated side of the membrane as shown in figure \(\PageIndex{1}\). The flow of solvent across the semipermeable membrane is called osmosis. Consider the osmotic pressure (\( \Pi \)) that develops on the more concentrated side of the membrane due to solvent molecules moving from the dilute to the concentrated side of the membrane. This situation illustrated in figure \(\PageIndex{1}\).

As the pressure builds up on the high concentration side, the solvent level will rise by a height, \(h\). The hydrostatic pressure is the difference between the pressure on the two sides of the semipermeable membrane and is given as
\[\Pi = \rho g h \label{EQ:hydrostatic} \]
where \(\rho \) is the solvent density, and \(g\) is the acceleration due to gravity. For an ideal solution, the osmotic pressure, \(\Pi \), resembles the form of the ideal gas law:
\[\Pi V = n RT \label{EQ:osmotic1} \]
Using the fact that \( n/V \) is the concentration (i.e. number of moles per unit volume of solution), we can rewrite Equation \ref{EQ:osmotic1} in terms of the molarity (M):
\[\Pi = \underline{M} RT \label{EQ:osmotic2} \]
If we define the mass concentration (C) of the solute, then we can rewrite \ref{EQ:osmotic2} as:
\[\Pi = \frac{CRT}{M_w} \implies \frac{\Pi}{RT C} = \frac{1}{M_w} \label{EQ:osmotic3} \]
where \(M_w \) is the molar mass of the solute molecule, and \(C\) is the mass concentration of the solute (\( C = n M_w /V \)). In this case we can extract the molar mass as:
\[M_w = \frac{RTC}{\Pi}\nonumber\]
from any single measurement of the osmotic pressure.
Equation \ref{EQ:osmotic3} assumes we are dealing with an ideal solution in which the solute particles are non-interacting. If this is not the case than a plot of \(\Pi\) versus C will not be linear as implied by equation \(\ref{EQ:osmotic3}\). Instead, if we want to consider a non-ideal solution of weakly interacting molecules, we can expand Equation \ref{EQ:osmotic3} in powers of \(C \) to obtain a virial equation of state for the osmotic pressure of a real solution:
\[\frac{\Pi}{RT C} = \frac{1}{M_w} \left[ 1 + B'_2 C + B'_3 C^2 + B'_4 C^3 + . . . \right] \label{EQ:virial_osmotic} \]
where \( B'_2, B'_3, B'_4 ...\) are the second, third, and fourth osmotic virial coefficients.
In the dilute limit (\(C\) << 1), we can truncate the expansion at the second virial coeffient:
\[\frac{\Pi}{RT C} = \frac{1}{M_w} \left[ 1 + B'_2 C \right] \label{EQ:virial_osmotic2} \]
The second virial coefficient \(B' \) is related to the interactions between particles. The second virial coefficent may be positive or negative depending on the nature of the particle-particle interactions. A negative coefficient corresponds to net attractive interactions, and a positive coefficient correspond to repulsive interactions.
Equation \(\ref{EQ:virial_osmotic2}\) is sometimes used to estimate the molar mass of macromolecules. Consider the following data for a hypothetical macromolecule that cannot pass through the membrane. Experimentally you measure the mass concentration of the macromolecule, the solution density and the height the solution rises to counteract the osmotic pressure between the side with no macromolecules and the side containing macromolecules.
Table \(\PageIndex{2}\): mass concentration, solution density and height from osmotic pressure for unknown macromolecule at 298.0 K.
| C (mg/mL) | \(\rho\) (g/mL) | h (cm) |
| 0.531 | 1.005 | 1.10 |
| 1.678 | 1.017 | 5.38 |
| 2.444 | 1.024 | 9.65 |
| 3.204 | 1.032 | 14.97 |
| 4.113 | 1.041 | 22.72 |
Solution
The linearized form of equation \(\ref{EQ:virial_osmotic2}\) suggests that the intercept of a plot of \(\frac{\Pi}{RTC}\) is the inverse of the molecular weight. Alternatively, with modern fitting software we could fit \(\Pi\) versus C which would follow the form:
\[\Pi = \frac{RT}{M_w}C + \frac{RTB_2}{M_w}C^2 = aC + bC^2\nonumber\]
Thus, Mw = RT/a. This gives the same value as calculating \(\frac{\Pi}{RTC}\) making the plot followed by taking the inverse of the intercept. The more traditional method is illustrated immediately below:
1) Calculate values for \(\Pi\) using \(\Pi = \rho g h\), equation \(\ref{EQ:hydrostatic}\). It is best to decide on units. For this example everything is converted to SI. So C is converted to kg/m3 by multiplication by (kg/106 mg)(106mL/m3) = 1(kg/m3)/(mg/mL) and cm are converted to meters.
2) Calculated the reduced osmotic pressure \(\frac{\Pi}{RTC}\).
Results of 1 and 2 are shown in the table below:
| C (mg/mL) | \(\Pi\) (Pa) | \(\frac{\Pi}{RTC}\) (mol/kg) |
| 0.531 | 108 | 0.0824 |
| 1.678 | 537 | 0.129 |
| 2.444 | 969 | 0.160 |
| 3.204 | 1515 | 0.1908 |
| 4.113 | 2319 | 0.2276 |
3) Plot and fit.

4) Calculate Mw as the inverse of the intercept.
\[M_w = \frac{1}{0.06096\, mol/kg} = 16.40 \,kg/mol\nonumber\]
A plot of \(\Pi\) versus concentration shows a definite positive curvature. So, this analysis is necessary to account for non-ideality.
Osmotic pressure plays an important role in biology. For example, trees use osmotic pressure to transport water from the roots to the upper branches. The effect of osmotic pressure on the cell is illustrated in figure \(\PageIndex{2}\). When red blood cells are placed in a salt solution having a lower concentration than the intracellular fluid, the solution is hypotonic, and the cell will gain water through osmosis in an attempt to equalize the osmotic pressure. This situation is illustrated in figure \(\PageIndex{2}\) (a). The cells will swell and potentially burst. When red blood cells are placed in a salt solution with the same osmotic pressure as the intracellular fluid, the solution is isotonic with respect to the cytoplasm. This situation is illustrated in See figure \(\PageIndex{2}\) (b). Finally, when red blood cells are placed in a solution with a higher salt concentration than the intracellular fluid, the solution is hypertonic and water inside the cell flows outside the cell in an attempt to equalize the osmotic pressure, causing the cell to shrink. This is the situation illustrated in figure \(\PageIndex{2}\) (c).

Derivation of osmotic pressure equation form
Osmosis is a process by which solvent can pass through a semi-permeable membrane (a membrane through which solvent can pass, but not solute) from an area of low solute concentration to a region of high solute concentration. The osmotic pressure is the pressure that when exerted on the region of high solute concentration will halt the process of osmosis.
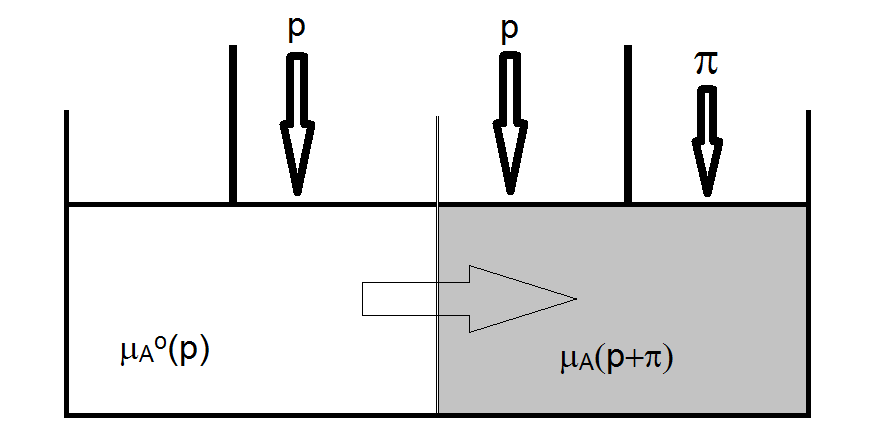
The nature of osmosis and the magnitude of the osmotic pressure can be understood by examining the chemical potential of a pure solvent and that of the solvent in a solution. The chemical potential of the solvent in the solution (before any extra pressure is applied) is given by
\[ \mu_A = \mu_A^o + RT \ln x_A \nonumber \]
And since xA < 1, the chemical potential is of the solvent in a solution is always lower than that of the pure solvent. So, to prevent osmosis from occurring, something needs to be done to raise the chemical potential of the solvent in the solution. This can be accomplished by applying pressure to the solution. Specifically, the process of osmosis will stop when the chemical potential solvent in the solution is increased to the point of being equal to that of the pure solvent. The criterion, therefore, for osmosis to cease is
\[ \mu_A^o(p) = \mu_A(x_b, +\Pi) \nonumber \]
To solve the problem to determine the magnitude of p, the pressure dependence of the chemical potential is needed in addition to understanding the effect the solute has on lowering the chemical potential of the solvent in the solution. The magnitude, therefore, of the increase in chemical potential due to the application of excess pressure p must be equal to the magnitude of the reduction of chemical potential by the reduced mole fraction of the solvent in the solution. We already know that the chemical potential of the solvent in the solution is reduced by an amount given by
\[ \mu^o_A - \mu_A = RT \ln x_A \nonumber \]
And the increase in chemical potential due to the application of excess pressure is given by
\[ \mu(p+\Pi) = \mu(p) + \int _{p}^{\Pi} \left( \dfrac{\partial \mu}{\partial p} \right)_T dp \nonumber \]
The integrals on the right can be evaluated by recognizing
\[\left( \dfrac{\partial \mu}{\partial p} \right)_T = V \nonumber \]
where \(V\) is the molar volume of the substance. Combining these expressions results in
\[ -RT \ln x_A = \int_{p}^{p+\Pi} V\,dp \nonumber \]
If the molar volume of the solvent is independent of pressure (has a very small value of \(\kappa_T\), constant temperature compressibility – which is the case for most liquids) the term on the right becomes.
\[ \int_{p}^{\Pi} V\,dP = \left. V p \right |_{p}^{p+\Pi} = V\Pi \nonumber \]
Also, for values of \(x_A\) very close to 1
\[ \ln x_A \approx -(1- x_A) = - x_B \nonumber \]
So, for dilute solutions
\[ x_B RT = V\Pi \nonumber \]
Or after rearrangement
\[ \Pi = \dfrac{x_B RT}{V} \nonumber \]
again, where \(V\) is the molar volume of the solvent. And finally, since \(x_B/V\) is the concentration of the solute \(B\) for cases where \(n_B \ll n_A\). This allows one to write a simplified version of the expression which can be used in the case of very dilute solutions
\[ \Pi = [B]RT \nonumber \]
When a pressure exceeding the osmotic pressure \(\Pi\) is applied to the solution, the chemical potential of the solvent in the solution can be made to exceed that of the pure solvent on the other side of the membrane, causing reverse osmosis to occur. This is a very effective method, for example, for recovering pure water from a mixture such as a salt/water solution.
Colligative properties are properties that depend on the number of particles rather than their total mass. This implies that these properties can be used to measure molar mass. Colligative properties include:
- melting point depression
- boiling point elevation
- osmotic pressure
Problems
Problem \(\PageIndex{1}\). The osmotic pressure of a protein in solution at 298 K (under crystallization conditions) was measured at the following concentrations:
| Concentration (g L-1) | osmotic pressure (10-2 kPa) |
| 0.50 | 1.85 |
| 1.00 | 3.68 |
| 1.50 | 5.48 |
| 2.00 | 7.25 |
| 2.50 | 9.00 |
Assuming that the osmotic virial equation of state can be truncated after the second virial coefficient,
a) What is the molar mass of the protein?
b) What is the value of the second osmotic virial coefficient in units of L/g?
c) Based on the sign of the second virial coefficient, what might you speculate about the average contribution of intermolecular interactions under crystallization conditions? (For reference see: A. George and W.W. Wilson, "Predicting Protein Crystallization from Dilute Solution Property," Acta Cryst. (1994). D50, 361-365).