
1.7: Phase Equilibria and Mixtures
- Page ID
- 512047
This page is a draft and is under active development.

\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\( \newcommand{\dsum}{\displaystyle\sum\limits} \)
\( \newcommand{\dint}{\displaystyle\int\limits} \)
\( \newcommand{\dlim}{\displaystyle\lim\limits} \)
\( \newcommand{\id}{\mathrm{id}}\) \( \newcommand{\Span}{\mathrm{span}}\)
( \newcommand{\kernel}{\mathrm{null}\,}\) \( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\) \( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\) \( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\id}{\mathrm{id}}\)
\( \newcommand{\Span}{\mathrm{span}}\)
\( \newcommand{\kernel}{\mathrm{null}\,}\)
\( \newcommand{\range}{\mathrm{range}\,}\)
\( \newcommand{\RealPart}{\mathrm{Re}}\)
\( \newcommand{\ImaginaryPart}{\mathrm{Im}}\)
\( \newcommand{\Argument}{\mathrm{Arg}}\)
\( \newcommand{\norm}[1]{\| #1 \|}\)
\( \newcommand{\inner}[2]{\langle #1, #2 \rangle}\)
\( \newcommand{\Span}{\mathrm{span}}\) \( \newcommand{\AA}{\unicode[.8,0]{x212B}}\)
\( \newcommand{\vectorA}[1]{\vec{#1}} % arrow\)
\( \newcommand{\vectorAt}[1]{\vec{\text{#1}}} % arrow\)
\( \newcommand{\vectorB}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\( \newcommand{\vectorC}[1]{\textbf{#1}} \)
\( \newcommand{\vectorD}[1]{\overrightarrow{#1}} \)
\( \newcommand{\vectorDt}[1]{\overrightarrow{\text{#1}}} \)
\( \newcommand{\vectE}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash{\mathbf {#1}}}} \)
\( \newcommand{\vecs}[1]{\overset { \scriptstyle \rightharpoonup} {\mathbf{#1}} } \)
\(\newcommand{\longvect}{\overrightarrow}\)
\( \newcommand{\vecd}[1]{\overset{-\!-\!\rightharpoonup}{\vphantom{a}\smash {#1}}} \)
\(\newcommand{\avec}{\mathbf a}\) \(\newcommand{\bvec}{\mathbf b}\) \(\newcommand{\cvec}{\mathbf c}\) \(\newcommand{\dvec}{\mathbf d}\) \(\newcommand{\dtil}{\widetilde{\mathbf d}}\) \(\newcommand{\evec}{\mathbf e}\) \(\newcommand{\fvec}{\mathbf f}\) \(\newcommand{\nvec}{\mathbf n}\) \(\newcommand{\pvec}{\mathbf p}\) \(\newcommand{\qvec}{\mathbf q}\) \(\newcommand{\svec}{\mathbf s}\) \(\newcommand{\tvec}{\mathbf t}\) \(\newcommand{\uvec}{\mathbf u}\) \(\newcommand{\vvec}{\mathbf v}\) \(\newcommand{\wvec}{\mathbf w}\) \(\newcommand{\xvec}{\mathbf x}\) \(\newcommand{\yvec}{\mathbf y}\) \(\newcommand{\zvec}{\mathbf z}\) \(\newcommand{\rvec}{\mathbf r}\) \(\newcommand{\mvec}{\mathbf m}\) \(\newcommand{\zerovec}{\mathbf 0}\) \(\newcommand{\onevec}{\mathbf 1}\) \(\newcommand{\real}{\mathbb R}\) \(\newcommand{\twovec}[2]{\left[\begin{array}{r}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\ctwovec}[2]{\left[\begin{array}{c}#1 \\ #2 \end{array}\right]}\) \(\newcommand{\threevec}[3]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\cthreevec}[3]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \end{array}\right]}\) \(\newcommand{\fourvec}[4]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\cfourvec}[4]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \end{array}\right]}\) \(\newcommand{\fivevec}[5]{\left[\begin{array}{r}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\cfivevec}[5]{\left[\begin{array}{c}#1 \\ #2 \\ #3 \\ #4 \\ #5 \\ \end{array}\right]}\) \(\newcommand{\mattwo}[4]{\left[\begin{array}{rr}#1 \amp #2 \\ #3 \amp #4 \\ \end{array}\right]}\) \(\newcommand{\laspan}[1]{\text{Span}\{#1\}}\) \(\newcommand{\bcal}{\cal B}\) \(\newcommand{\ccal}{\cal C}\) \(\newcommand{\scal}{\cal S}\) \(\newcommand{\wcal}{\cal W}\) \(\newcommand{\ecal}{\cal E}\) \(\newcommand{\coords}[2]{\left\{#1\right\}_{#2}}\) \(\newcommand{\gray}[1]{\color{gray}{#1}}\) \(\newcommand{\lgray}[1]{\color{lightgray}{#1}}\) \(\newcommand{\rank}{\operatorname{rank}}\) \(\newcommand{\row}{\text{Row}}\) \(\newcommand{\col}{\text{Col}}\) \(\renewcommand{\row}{\text{Row}}\) \(\newcommand{\nul}{\text{Nul}}\) \(\newcommand{\var}{\text{Var}}\) \(\newcommand{\corr}{\text{corr}}\) \(\newcommand{\len}[1]{\left|#1\right|}\) \(\newcommand{\bbar}{\overline{\bvec}}\) \(\newcommand{\bhat}{\widehat{\bvec}}\) \(\newcommand{\bperp}{\bvec^\perp}\) \(\newcommand{\xhat}{\widehat{\xvec}}\) \(\newcommand{\vhat}{\widehat{\vvec}}\) \(\newcommand{\uhat}{\widehat{\uvec}}\) \(\newcommand{\what}{\widehat{\wvec}}\) \(\newcommand{\Sighat}{\widehat{\Sigma}}\) \(\newcommand{\lt}{<}\) \(\newcommand{\gt}{>}\) \(\newcommand{\amp}{&}\) \(\definecolor{fillinmathshade}{gray}{0.9}\)Based on "Equilibria in Biochemical Systems" by Serge L. Smirnov and James McCarty, LibreTexts is licensed under CC BY-NC-SA .
In this chapter we extend the concept of the Gibbs energy to phase transitions and mixtures. In the case of mixtures, the number of moles of the different components can change as a result of a chemical reaction or a phase transition. The partial molar Gibbs energy or chemical potential can be used to determine the spontaneity a process. We first derive an expression for the chemical potential. Then we will consider the impacts of pressure and temperature and use that to understand phase transitions, vapor pressure of volatile liquids, and ideal solutions. We then introduce the concept of the activity to write a general expression for the chemical potential.
- Know the definition of the chemical potential as the partial molar Gibbs energy, and be able to analyze the spontaneity of a phase transition based on the change in chemical potential.
- Be able to use the chemical potential to calculate the change in Gibbs energy for a process involving changing number of moles.
- Understand how to model the shapes of the phase boundaries in a pressure versus temperature phase diagram.
- Understand the definition of the activity and how it can be used to describe both ideal and real solutions.
Gibbs energy and phase equilibria
For a phase transition in equilibrium at the phase transition temperature, such as the freezing of liquid water at 0 °C, the process is reversible. At equilibrium, the change in the molar Gibbs energy \(\Delta G_{molar}= 0\), meaning that if two phases are at equilibrium,
\[G_{molar} (solid) = G_{molar} (liquid)\]
Notice here that the molar Gibbs energy is the same for the two phases. The molar Gibbs energy is an intensive variable (Gibbs energy per mole). We must use the molar Gibbs energy because the phase equilibrium is independent of the amount of substance. For example, we could have a small ice cube in equilibrium with a large volume of water at 0 °C.
If we have multiple species in our system, the intensive variable of interest is the partial molar Gibbs energy that is defined for the ith component of the system as:
\[G_{molar, i}=\left(\frac{\partial{G}}{\partial{n_i}}\right)_{T,P,n_j}\]
where \(n_i\) is the number of moles of the ith component, and \(n_j\) is the number of moles of all the other components in the system. The total Gibbs energy is a function of the number of moles of each species:
\[G=\sum_i{n_i G_{molar,i}}\]
The partial molar Gibbs, \(G_{molar, i}\) also shows how infinitesimal changes in the Gibbs energy, dG, depend on infinitesimal changes in the number of moles of a component (n1, n2, …):
\[dG = G_{molar, 1} dn_1 + G_{molar, 2} dn_2 + …\label{equation:dG}\]
The chemical potential
Because the partial molar Gibbs is used so often when considering equilibria, we give this quantity a special name called the chemical potential which gets the Greek symbol μ, and we write the chemical potential of the ith component as:
\[μ_i=G_{molar, i}=\left(\frac{\partial{G}}{\partial{n_i}}\right)_{T,P,n_j}\]
Substituting into Equation \(\ref{equation:dG}\) gives:
\[dG = \mu_1 dn_1 + \mu_2 dn_2 + … = \sum_i{\mu_i dn_i}\label{equation:dGfromdmu}\]
Consider the spontaneous transfer of some moles of a molecule from state A to state B as shown in figure 1.7.1 .
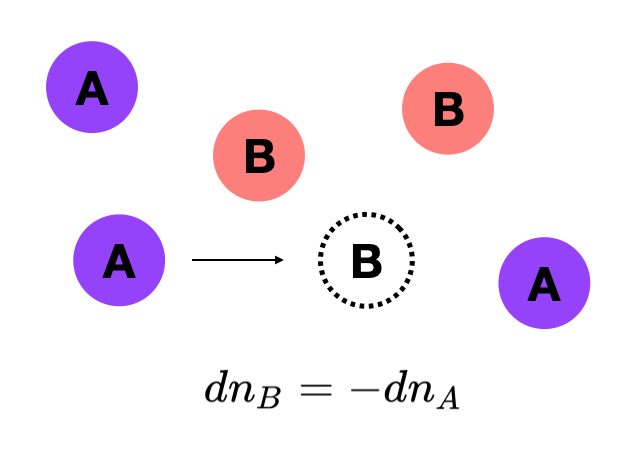
The change in the number of moles in state A, dnA will be equal and opposite to the change in the number of moles in state B, dnB, so we can write the total change in Gibbs energy from equation \(\ref{equation:dGfromdmu}\) as
\[dG = \mu_Adn_A+\mu_Bdn_B = (\mu_B−\mu_A)dn_B\]
We now ask ourselves, when will the transition from state A into state B become spontaneous? For a spontaneous process dG < 0. Thus, the transition of dnB moles from state A to state B, will be spontaneous if
\[(\mu_B−\mu_A)dn_B \lt 0 \implies \mu_B \lt μ_A\]
We see that matter flows in the direction of lower chemical potential.
Key Result: Matter flows spontaneously from high chemical potential to low chemical potential. The flow of matter will continue until the chemical potentials are equal, which is the equilibrium condition.
How does changes in conditions change G and thus \(\mu\)?
In order to determine how the equilibria between phases change as conditions change we need to understand the dependence of G (and thus \(\mu\)) on state variables such as T and P.
Changes in pressure
Recall that:
\[\left(\frac{\partial{G}}{\partial{P}}\right)_T = V \label{EQ:dGdPconstT}\]
or for \(\mu\):
\[\left(\frac{\partial{\mu}}{\partial{P}}\right)_T = V_{molar} \label{EQ:dmudPconstT}\]
For an ideal gas Vmolar depends on P :
\[\left(\frac{\partial{\mu}}{\partial{P}}\right)_T = \frac{RT}{P} \label{EQ:dmudPconstTIdealGas}\]
For incompressible liquids and solids Vmolar is a constant.
Changes in temperature
Recall that the fundamental differential relation for free energy is:
\[dG = VdP - SdT \implies d\mu = V_{molar}dP - S_{molar}dT \label{EQ:dGdiff}\]
If pressure is constant dP = 0 and and equation \(\ref{EQ:dGdiff}\) becomes:
\[d\mu = - S_{molar}dT \implies \left( \frac{ \partial{\mu}}{ \partial{T}}\right)_P = -S_{molar} \label{EQ:dmudiffconstP}\]
Single species phase equilibria
The simplest use of these relations is to understand the shape of the curves in a pressure versus temperature phase diagram for a pure substance. If we have two phases A and B equilibrium is when \(\mu_A = \mu_B\). We need to understand how the combinations of P and T where this is true change.
Impact of changes in P
First, we consider how P impacts the values of \(\mu\) at constant T. Since matter takes up space \(V_{molar} \gt 0\) (equations \(\ref{EQ:dmudPconstT}\) and \(\ref{EQ:dmudPconstTIdealGas}\)), \(\mu\) increases with pressure (P). The slope of the increase is the constant \(V_{molar} \) for solids and liquids, while for gases the slope is steepest at lower pressures. Without knowing actual values of \(\mu\) we can sketch a \(\mu\) versus P diagram showing when the chemical potential of pairs of phases are equal. At high pressure the most stable phase will be the one with the lowest \(V_{molar} \) (highest density, usually the solid phase) because as pressure increases that one will increase the least. In figure \(\PageIndex{2}\) this is indicated by the right most point on the line for \(\mu_s\). Likewise the liquid phase will have a slightly higher \(\mu_l\) indicated by the right most point on the line for \(\mu_l\) in figure \(\PageIndex{2}\). The gas phase has the highest \(\mu_g\) at high pressure, with a \(\mu\) that drops off more steeply as the pressure drops.

Figure \(\PageIndex{2}\): Variation of \(\mu\) for solids, liquids and gases versus pressure. Each phase is stable in the region where \(\mu\) of that phase is lowest. In this example only a little sliver near P = 0 favors the gas phase.
As seen in figure \(\PageIndex{2}\), this leads to the lowest pressure equilibrium occurring where the gas and liquid lines cross. The next equilibrium occurs at a slightly higher pressure where the gas and solid lines cross. The final equilibrium occurs at the highest pressure between the liquid and solid. As noted matter flows towards the lowest \(\mu\). Thus figure \(\PageIndex{2}\) can be divided into three pressure regions demarcated by the pressures of the equlibriums: 1) the lowest pressure region is where the gas phase is thermodynamically favored; 2) the middle pressure region where the liquid phase is favored; and 3) the high pressure region where the solid phase is favored. Thus, if we started with a sample in the gas phase and increased the pressure it would undergo a phase transition to the liquid phase at the left edge of the liquid region. The chemical potential \(\mu\) would then follow the liquid line until reaching the right edge of the liquid region. The material then undergoes a phase transition to the solid phase and follows the solid \(\mu\) line as the pressure increases more.
Impact of changes in T
Second, we consider how T impacts the values of \(\mu\) at constant P. Since entropy is always greater than zero the slope of \(\mu\) (equation \(\ref{EQ:dmudiffconstP}\)) is negative for all phases. As before, we can draw a qualitative diagram showing the relations of the chemical potentials as the temperature is varied. At any temperature \(S_{molar}(g) \gt S_{molar}(l) \gt S_{molar}(s) \). Thus at low temperatures the most stable phase will be the the solid; the next most stable the liquid and the least stable the gas. This relationship is indicated by the initial (leftmost) values of the \(\mu\) for each phase in figure \(\PageIndex{3}\). Although entropy increases with temperature, so that the slopes of the curves should get more steeply negative, in this diagram we assume the temperature range is small enough that \(S_{molar}\) is nearly constant leading to linear behavior over the temperature range plotted. Once again this leads to three places where two of the phases are in equilibrium and three regions of phase stability: 1) solid stable at low temperature; 2) liquid stable in an intermediate range and 3) gas stable at the highest temperatures (see figure \(\PageIndex{3}\)).

Figure \(\PageIndex{3}\): Variation of \(\mu\) for solids, liquids and gases versus temperature. Each phase is stable in the region where \(\mu\) of that phase is lowest. We hava assumed that the temperature range of this figure is small enough that the entropies of each phase is nearly constant, leading to a nearly constant slope.
Pressure versus temperature phase diagrams
Typical phase diagrams for pure substances display curves in P-T space, where the curves indicate the boundaries between the phases. Alternatively, you can think of the curves as representing the combinations of P and T where the neighboring phases A and B are in equilibrium (\(\mu_A = \mu_B\)). To describe the shapes of these curves we require expressions for \(\frac{dP}{dT}\) along these boundaries. First we will derive the generic relation and then examine the implications for the shape of each boundary.
Beginning with equation \(\ref{EQ:dGdiff}\) in its chemical potential form we note that chemical potentials of the two species must stay equal to stay on the boundary. This means the the changes in the two chemical potentials must be equal:
\[d\mu_A = V_{molar,A}dP - S_{molar,A}dT = d\mu_B = V_{molar,B}dP - S_{molar,B}dT \implies V_{molar,A}dP - S_{molar,A}dT =V_{molar,B}dP - S_{molar,B}dT \label{EQ:dmuA=dmuB}\]
Rearranging the last expression to get the molar volumes on one side and the molar entropies on the other leads to:
\[ S_{molar,B}dT - S_{molar,A}dT = V_{molar,B}dP - V_{molar,A}dP \implies (S_{molar,B} - S_{molar,A})dT = (V_{molar,B} - V_{molar,A})dP \label{EQ:DSdT=DVdP}\]
Notice that in equation \(\ref{EQ:DSdT=DVdP}\) the collected differences are just the changes in the molar entropy and volume on shifting phases. So this can be rewritten as:
\[ \Delta S_{molar}dT = \Delta V_{molar}dP \label{EQ:DSdT=DVdP2}\]
Dividing both sides by \(\Delta V_{molar}\) and dT yields an expression for the change in P for a change in T:
\[ \frac{dP}{dT} = \frac{\Delta S_{molar}}{\Delta V_{molar}} = \frac{\Delta S_{trs}}{\Delta V_{trs}}\label{EQ:Clapyron}\]
where in the final version the subscript 'trs' stands for transition ('vap' for vaporization, 'fus' for melting, 'sub' for sublimation). This is the fundamental form of the Clapeyron equation. A very useful alternative form of this equation makes use of the fact that when P and T are constant (at a particular equilibrium point on the boundary) \(\Delta S_{trs} = \frac{\Delta H_{trs}}{T_{trs}}\). Substituting this into equation \(\ref{EQ:Clapyron}\) yields:
\[ \frac{dP}{dT} = \frac{\Delta H_{trs}}{T_{trs}\Delta V_{trs}}\label{EQ:Clapyron2}\]
With this expression we can estimate the slope at any point along a phase boundary given enthalpy, temperature and molar volume change or a way to estimate them.
Carbon dioxide exhibits a phase diagram (figure \(\PageIndex{4}\)) that is typical of substances that exist as solids, liquids and gases between a 0 and a few hundred atmospheres of pressure and between temperatures that are easily achieved in the lab. Thus it is a good example to keep in mind as we look at the slopes of the phase boundaries.

Figure \(\PageIndex{4}\) Phase diagram of CO2. This is an example of a typical phase diagram.
Solid — liquid phase boundary
In most cases Vmolar(solid) < Vmolar(liquid) so \(\Delta V_{trs}\gt 0\). Also \(\Delta H_{trs}\gt 0\). Since temperature is also positive, equation \(\ref{EQ:Clapyron2}\) implies that the solid - liquid coexistence boundary will have a positive slope. As the enthalpy and volume change are nearly constant with pressure and temperature the slope will slowly decrease as the temperature increases. In the part of the CO2 phase diagram shown in figure \(\PageIndex{4}\) this phase boundary appears to grow linearly. This is because the ratio \(\frac{\Delta H}{\Delta V}\) is so large that the temperature changes little in the pressure range considered. We can rearrange \(\ref{EQ:Clapyron2}\) and integrate to get an expression for the change in P for a given change in T assuming \(\frac{\Delta H}{\Delta V}\) is constant:
\[\frac{dP}{dT} = \frac{\Delta H}{T \Delta V} \implies dP = \frac{\Delta H}{T \Delta V}dT \]
so, assuming constant enthalpy and volume changes:
\[ \int_{P_1}^{P_2}dP = \int_{T_1}^{T_2}\frac{\Delta H}{T \Delta V}dT \implies \Delta P = \frac{\Delta H}{\Delta V} \int_{T_1}^{T_2}\frac{dT}{T}\]
which integrates to:
\[\Delta P = \frac{\Delta H}{\Delta V} ln \left(\frac{T_2}{T_1}\right)\]
So given a starting point on the curve we can estimate nearby P and T combinations on the curve.
Note that water has a solid - liquid phase boundary with a negative slope because Vmolar(ice) > Vmolar(water); thus \(\Delta V \lt 0\) (see figure \(\PageIndex{5}\)).

Figure \(\PageIndex{5}\): Two Versions of the Phase Diagram of Water. (a) In this graph with linear temperature and pressure axes, the boundary between ice and liquid water is almost vertical. (b) This graph with an expanded scale illustrates the decrease in melting point with increasing pressure.
Liquid — gas and solid — gas phase boundaries
In this case Vmolar(g) is much greater than Vmolar(l) or Vmolar(s). For example near room temperature the molar volume of liquid water is about 18 mL, while the molar volume of gaseous water is about 800 L. So Vmolar(g) is around 40000 times larger than that of the liquid or solid. Thus, to within about 1 part in 105:
\[\Delta V_{trs} = V_{molar}(g) - V_{molar}(\text{s or l}) \approx V_{molar}(g)\label{EQ:DVtrsliqtogas}\]
Substituting this into the Clapeyron equation \(\ref{EQ:Clapyron2}\) and assuming the gas behaves ideally, \(V_{molar} = \frac{RT}{P}\), yields:
\[\frac{dP}{dT} = \frac{\Delta H_{trs}}{T_{trs}V_{molar}(g)} = \frac{P\Delta H_{trs}}{RT^2}\label{EQ:ClausClapyron}\]
Collecting all the Ps on one side and Ts on the other allows us to integrate one side versus P and the other versus T. First consider the indefinite integral:
\[\int\frac{dP}{P} = \int\frac{\Delta H_{trs}}{RT^2}dT \implies lnP = \frac{-\Delta H}{RT} + C\label{EQ:ClausClapyron2}\]
where we have collected the constants of integration from both sides into C. Exponentiating both sides and calling \(e^{C}\) C' gives us an expression for the temperature dependence of P along the phase boundary:
\[P = C^{\prime} exp \left(\frac{-\Delta H}{RT}\right) = C^{\prime} e^{\frac{-\Delta H}{RT}}\label{EQ:ClausClapyron3}\]
So, we expect to see exponentially increasing curves as T increases (\(\frac{-\Delta H}{RT} \to 0\)). These terminate at the triple and critical points, so do not blow up. We can also do the definite integrals of equation \(\ref{EQ:ClausClapyron}\), which yields the most commonly used form of the Clausius-Clapeyron equation:
\[ln \left(\frac{P_2}{P_1}\right) = \frac{-\Delta H}{R}\left(\frac{1}{T_2} - \frac{1}{T_1}\right)\label{EQ:ClausClapyron4}\]
Vapor pressure of a pure substance
Equations \(\ref{EQ:ClausClapyron3}\) and \(\ref{EQ:ClausClapyron4}\) also show how the vapor pressure of a pure substance varies with temperature.
NOTE: keep in mind that the assumption that \(\Delta H_{vap}\) is constant means that the equation is only valid over relatively small temperature changes. For larger changes the the variation must be taken into account, usually by measuring the values experimentally.
Mixtures
For a multi-component system of ideal gases, chemical potential for each component is related to the partial pressure Pi of each species compared to the standard potential at 1 bar. Rearranging and integrating equation \(\ref{EQ:dmudPconstTIdealGas}\) we get:
\[\int_{P^o}^{P_i}d\mu = \int_{P^o}^{P_i}\frac{RT}{P}dP \implies \Delta \mu = RT ln \left(\frac{P_i}{P^o}\right)\]
Adding this to the standard chemical potential of the species \(\mu^o_i\) yields:
\[\mu_i = \mu^o_i + RT ln \left(\frac{P_i}{P^o}\right)\label{EQ:mugas}\]
Key Result: For a mixture of ideal gases, the chemical potential of the ith species is \(\mu_i = \mu^o + RT ln \left(\frac{P_i}{P^o}\right)\) where Pi is the partial pressure of the gas and \(\mu^o_i\) is the standard chemical potential of component i when its partial pressure is 1 bar.
Thermodynamics of mixing volatile liquids
Figure \(\PageIndex{6}\) shows a pure liquid at equilibrium with its vapor in a closed container.

Since the system is at equilibrium the chemical potentials are equal:
\[\mu^*_{vapor} = \mu^*_{liquid}\label{EQ:muvolatile}\]
where the asterisk (*) indicates a pure substance. From equation \(\ref{EQ:mugas}\) for the gas phase we can write:
\[\mu^*_{vapor} = \mu^*_{liquid} = \mu^o_{vapor} + RT ln \left(\frac{P^*}{P^o}\right)\label{EQ:muliq}\]
where P∗ is the vapor pressure and \(\mu^o_{vapor}\) is the chemical potential at Po = 1 bar.
Now consider a mixture of volatile liquids as shown in figure \(\PageIndex{7}\)

Since both components are in equilibrium with their vapors, the chemical potential for each component is still equal in the two phases. For example, for component A, we have:
\[\mu_A (l) =\mu_A (g) =\mu^o_A (g) + RT ln \left(\frac{P_A}{P^o}\right)\label{EQ:muA}\]
where PA is the partial pressure of vapor A. Because, \(\mu^o_{vapor}=μ^o_A(g)\), we subtract equation \(\ref{EQ:muliq}\) from equation \(\ref{EQ:muA}\) to obtain:
\[\mu_A (l) – \mu^*_A (l) =RT \ln \left(\frac{P_A}{P^o}\right) -RT \ln \left(\frac{P^*}{P^o}\right) \implies \mu_A (l) = \mu^*_A (l) + RT \ln \left(\frac{P_A}{P_A^*}\right)\label{EQ:muA2}\]
Thus, from equation \(\ref{EQ:muA2}\), the chemical potential of a liquid in a mixture, μA(l), is given in terms of the chemical potential of the pure liquid (\(μ^∗_A(l)\)) and the ratio of the vapor pressure in the pure state over the vapor pressure in the mixture.
The relationship between the two vapor pressures in equation is given by Raoult’s law which states that the vapor pressure of a substance in a mixture is the product of its vapor pressure as a pure liquid and its mole fraction:
\[P_A = x_A P_A^*\label{EQ:Raoults}\]
where xA is the mole fraction of component A in the mixture. Inserting equation \(\ref{EQ:Raoults}\) into equation \(\ref{EQ:muA2}\) gives the final expression for the chemical potential of a liquid in an ideal mixture:
\[\mu_A (l) = \mu^*_A (l) + RT ln(x_A)\label{EQ:muLiqIdeal}\]
Raoult's law is based on a lifetime of work measuring vapor pressures (partial pressures) of liquid vapor above solutions by F. Raoult in the 1800s. It is only valid for solutions that behave ideally or are very dilute (xA ≈ 1).
Very dilute conditions are met when the volatile liquid of interest is in large excess (often the case for the solvent in a solution). The closer the mole fraction of the solvent is to 1, the more ideally it behaves. There are some cases where the two species being mixed are similar enough that ideal behavior is seen for all mole fractions. An example where ideal behavior is seen at all concentrations is when mixing benzene and methylbenzene.
Physical interpretation of Raoult's law: For an ideal mixture of two substances the fraction of each substance at the surface will be their mole fractions. This means that the chance a molecule of a substance can escape the surface will be reduced from the pure case by a factor of its mole fraction.
Thermodynamic interpretation of Raoult's law: A mixture has a higher entropy than a pure substance. This reduces the propensity of liquid molecules escaping into the gas phase to increase their entropy.
See Practice Problems \(\PageIndex{4}\) and \(\PageIndex{5}\).
Thermodynamics of ideal solutions
For the case of a solute dissolved in a solvent (liquid), the chemical potential of the solvent is the same as for a mixture of volatile liquids:
\[\mu_{solvent} (l) = \mu_{solvent}^* (l) + RT \ln x_{solvent}\]
For the case of the solute, it is often more convenient to express the chemical potential in terms of the molality m defined as
\[\text{molality} = \frac{\text{moles of solute}}{\text{mass of solvent in kg}} \nonumber\]
For the solute, the chemical potential is:
\[\mu_{solute} (l) = \mu_{solute}^{\circ}(l) + RT \ln \left( \frac{m_{solute}}{m^{\circ}}\right)\]
Note here the careful choice of the reference state for the solute. The references state is defined as a state of unit molality where mo = 1 mol kg-1.
Thermodynamics of real solutions
Under realistic conditions such as inside a cell or in a body of surface water (oceans and most lakes and streams) the conditions are far from that of an ideal solution. To account for more realistic conditions while maintaining the mathematical formulations as close to what we have derived as possible we write the chemical potential of a species as:
\[\mu_i = \mu^{\circ}_i + RT \ln a_i\]
where ai is called the activity and μ∘i is a reference state. For real solutions, the activity is given as
\[a_i = \gamma_i (m_i/m^{\circ})\label{EQ:mu_general}\]
where γi is called the activity coefficient that is a measure of the deviation from ideality. For an ideal solution, γ=1 and \(a_i = \frac{m_i}{m^o}\). Table \(\PageIndex{1}\) summarizes the expression of the activity and standard state for various substances. Note that for a pure sold and a pure liquid the activity is one.
| Substance | Standard State (μo) | activity (a) |
|---|---|---|
| solid | pure solid, 1 bar | 1 |
| liquid | pure liquid, 1 bar | 1 |
| gas | pure gas, 1 bar | P∗/(1 bar) |
| solvent | pure solvent | mole fraction xi |
| ideal solute | molality of 1 mol kg−1 | \(\frac{m_i}{m^o}\) |
| real solution | molality of 1 mol kg−1 | \(\gamma_i (m_i/m^{\circ})\) |
Practice Problems
Problem \(\PageIndex{1}\). At high pressure, graphite (density ρ=2.25 g/cm3) can be spontaneously converted into diamond (density ρ=3.51 g/cm3) through a solid-to-solid phase transition:
C (graphite) → C (diamond)
At 1 atm of pressure the standard molar Gibbs energy of this reaction is \(\Delta G^o\)=2.84 kJ/mol. At what pressure does the reaction become spontaneous (i.e. at what pressure does ΔGmolar=0)?
Problem \(\PageIndex{2}\). Consider the reversible freezing of liquid water into ice at a constant temperature of 0 °C and constant pressure of 1 atm.
H2O (l) ↔ H2O (g) 0 °C
Show that ΔG for this process is 0. (Hint: use the relation \(\Delta G = \Delta H – T\Delta S\)).
Problem \(\PageIndex{3}\). Which of the following has a higher chemical potential? (If neither, answer “same”)
(a) H2O (l) or H2O (s) at water’s normal melting point (0 ºC).
(b) H2O (l) or H2O (s) at -5 ºC and 1 bar.
Problem \(\PageIndex{4}\). Which would have the higher chemical potential? Benzene at 25 ºC and 1 bar or benzene in a 0.1 M toluene solution at 25 ºC and 1 bar.
Problem \(\PageIndex{5}\). Calculate the chemical potential of ethanol in solution relative to that of pure ethanol when its mole fraction is 0.40 at its boiling point (78.3 ºC.)